
Plan to attend the fourth
Montréal, Québec
Please join us at the Hotel Bonaventure Montreal as we host the fourth Canadian Symposium on Lysosomal Diseases (CSLD). We invite Physicians, Researchers, Registered Nurses, Genetic Counsellors and Trainees to attend this in-person Symposium.
The theme for this Symposium is
Canadian and international leaders and experts will present innovative trends and solution-focused best practices in the diagnosis and management of lysosomal diseases.
Time zone for these dates is Eastern Standard Time (EST)
29 October, 2026
30 October, 2026
12 October, 2026
13 – 14 November, 2026
900 De La Gauchetiere Ouest, Montréal, Québec
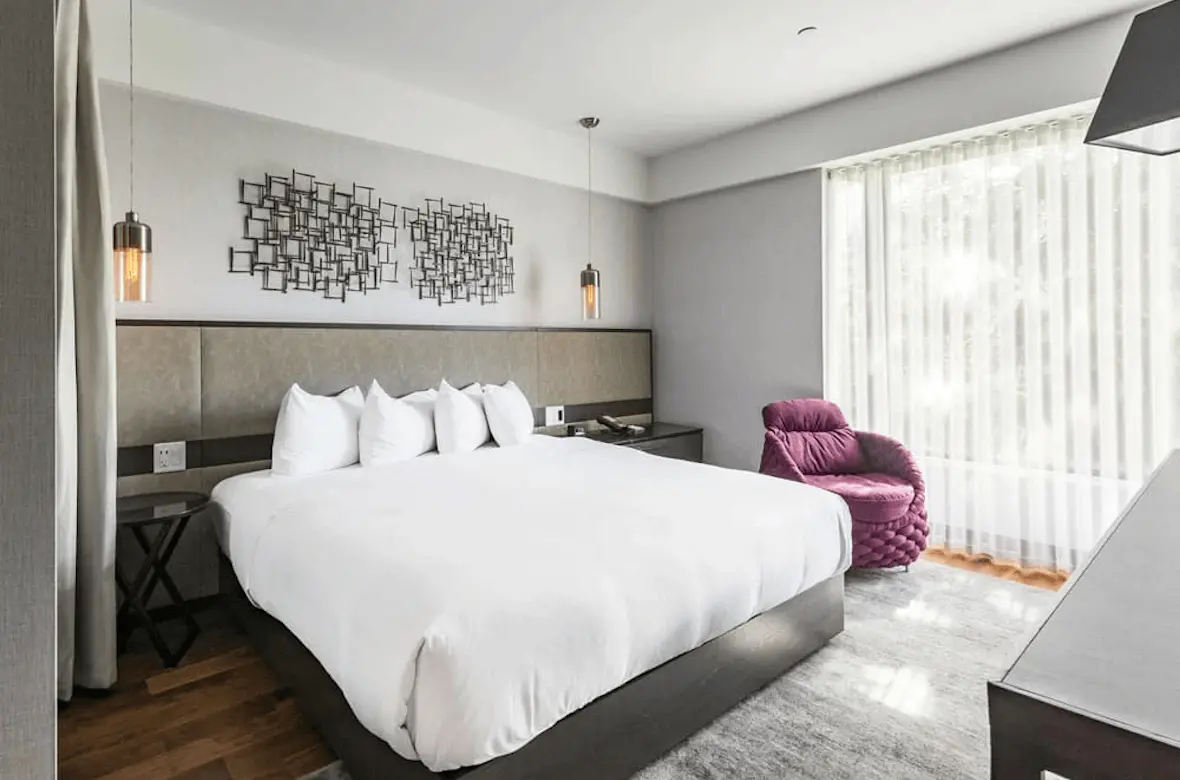


The newly renovated Bonaventure offers ideal facilities and technical support for the Fourth Canadian Symposium on Lysosomal Diseases (CSLD). The hotel features eco-friendly approaches to catering and event services.
The Hotel Bonaventure Montréal is the official hotel for the CSLD. The hotel features recently renovated guest rooms sure to please the most discriminating guests. The location is perfect—steps from the underground and a 10-minute walk to many major attractions, restaurants, historic sites and shopping. Among the many amenities are hot springs, a year-round outdoor pool and spa tub, salon/spa, complimentary WiFi, and an excellent in-house restaurant and bar.
You are responsible for booking your own accommodations.
Our Hotel Bonaventure block is now sold out.
All sessions take place in salle de bal de Montréal 1-2-3
Background: Enzyme replacement therapy (ERT) has transformed Fabry disease management. However, neutralizing anti-drug antibodies (ADAs), particularly in males with classic Fabry disease, can reduce effective enzyme exposure and substrate clearance, contributing to persistent lyso-Gb3 elevation and ongoing disease activity. Their detection does not always translate into clear therapeutic decisions.
Objective: To propose a practical framework for recognizing when neutralizing ADAs may contribute to inadequate treatment response and for guiding timely therapeutic adaptation.
Approach: We developed an action-oriented framework integrating ADA status, lyso-Gb3 response, clinical trajectory, and organ involvement. Persistent biochemical or clinical evidence of inadequate disease control should prompt consideration of neutralizing ADAs and reassessment of treatment strategy. Switching between agalsidase alfa and beta may not overcome clinically significant ADAs because of potential antibody cross-reactivity. Depending on genotype and clinical context, alternative approaches include pegunigalsidase alfa, migalastat for amenable GLA variants, and clinical trials of substrate-reduction or gene-based therapies. Immune-modulating strategies remain investigational, while optimization of organ-specific supportive care remains essential.
Conclusion: Neutralizing ADAs should not be viewed simply as a laboratory finding to monitor passively. In the appropriate clinical context, they may represent a clinically relevant mechanism of inadequate treatment response and should prompt timely reassessment of disease control and therapeutic strategy. The key is to integrate antibody status with biochemical response, clinical trajectory, and organ involvement to determine whether ADAs are compromising treatment and whether a change in therapeutic strategy is warranted.
Pegunigalsidase alfa (PA) is a new version of enzyme replacement therapy (ERT) for Fabry disease (FD) available in Canada. We report our early experience with this therapy. There are 8 adults with FD in Canada (50%F) mean age 43.8 who have received therapy with PA. They include 6 with missense variants, 2 with intronic splice site variants, 6 of classic phenotype and 2 of non-classsic renal/cardiac disease. Three patients were in PA clinical trials for 11-12 years; the other 5 have received PA for 1-13 months. Indications for ERT were pain (3), hearing loss (1), renal disease (2), strokes (2) and cardiac disease (3). Prior therapy included agalsidase alfa (6 )(mean 54.8 months) and migalastat (1); 1 pt was treatment naive. Plasma lysoGb3 and urine Gb3 fell in all. PA was well tolerated in most. The ERT nave patient developed infusion associated reactions with chills. This was controlled with acetaminophen, antihistamine, prednisone and slowing of the infusion rate. Antidrug antibody (ADA) tests i showed IgE in 1 but no anaphylaxis and non-neutralizing IgG in 2 (1/12,800; 1/100). Benefits included clearance of cornea verticillata, increased sweating, increased energy, slowing of eGFR decline, lessening of diarrhea and abdominal pain, and stabilization of cardiac disease in some patients. In conclusion, pegunigalsidase alfa is well tolerated in general and should meet the needs of some adults with FD who require higher dose ERT. More real world data are required as to prevalence of IgE and IgG ADA, target epitopes, associated reactions and impact on patient outcomes.
Background: MPS IIIA (Sanfilippo type A) is a progressive neurodegenerative disease characterized by developmental arrest, regression, and early death. UX111 (rebisufligene etisparvovec) is an intravenously delivered AAV9 vector encoding SGSH under investigation for MPS IIIA (PDUFA 19SEP2026).
Methods: Data were analyzed across two open-label studies (NCT02716246, NCT04088734) and an ongoing long-term follow-up (NCT04360265). Children received a single intravenous infusion of UX111. The Efficacy set included those who received 3x10^13 vg/kg UX111. The mITT set included a subset 2 years old or >2 years old with a BSID-III cognitive developmental quotient 60. CSF HS exposure was defined as time-normalized AUC of percentage change from baseline. Data snapshot: 22Sept2025.
Results: Median (min-max) follow-up was 4.8 (0.6, 8.5) years (n=33). Median (95% CI) CSF HS exposure was ‑63.98%(‑72.27, ‑54.84;p<0.0001)for the Efficacy set (n=27) and‑62.58%(‑74.26, -56.29;p<0.0001)for mITT (n=17).For mITT, LS mean change in model-estimated BSID-III raw scores from 24-60 months of age significantly improved vs Natural History for cognitive (23.2 points, p<0.0001), receptive communication (8.1 points, p=0.0076), expressive communication (11.1 points, p=0.0008), and fine motor (9.0 points, p=0.0026). Median (95% CI) annualized percentage change from baseline in total cortical brain volume was 0.83 (3.02, 6.75) in mITT and 8.04% (10.18, 5.46) in the Natural History set (p=0.0003). Children with more advanced disease at treatment (n=10) maintained key communication, ambulation, and/or eating/self-feeding skills.
Most UX111-related TEAEs were mild-to-moderate hepatic reactions that resolved spontaneously.
Conclusions: UX111 showed a positive treatment effect and manageable safety in MPS IIIA.
Sialidosis is a lysosomal storage disorder (LSD) caused by deficiency of neuraminidase 1 (NEU1), a sialidase localized primarily to lysosomes and the plasma membrane. NEU1 removes terminal sialic acid residues from glycoproteins and glycolipids. The impact of lysosomal dysfunction on ER stress has been demonstrated in several LSD models. In fact, our group has shown that ER stress is associated with disease progression in Sandhoff disease (SD) and that treatment with the chemical chaperone 4-phenylbutyric acid reduced ER stress, improved motor function, and extended lifespan, emphasizing the role of ER stress in LSD pathophysiology. To extend our studies to other LSDs, we investigated whether NEU1 deficiency results in ER stress in the mouse liver. Quantitative immunofluorescence and Western blot analyses were performed using liver tissues with complete (knockout) or partial (heterozygous) NEU1 deficiency. We found that both complete and partial NEU1 deficiency significantly altered the expression and subcellular localization of the ER stress-associated proteins GRP78, KDEL, and GRP94, as well as the lipid droplet-associated protein PLIN2. Notably, the magnitude and pattern of these alterations differed between the homozygous and heterozygous mice, indicating that hepatic ER stress responses are influenced by the extent of NEU1 deficiency. Collectively, these findings identify an association between NEU1 deficiency, ER homeostasis, and hepatic lipid metabolism, and provide new insight into the molecular interplay between lysosomal dysfunction and ER stress in NEU1-associated disease.
Background: Pompe disease is a rare, inherited genetic disease caused by mutations altering GAA gene functions. It leads to lysosomal glycogen accumulation in cells/tissues (particularly, skeletal muscles). Pompe disease (PD) involves energy-source shifts, mitochondrial dysfunction, and disrupted autophagy, which may affect the circulating lipidome.
Objectives: 1) To identify novel biomarkers in plasma samples from untreated Infantile-Onset Pompe Disease (IOPD) and Late-Onset Pompe Disease (LOPD) patients using semi-targeted lipidomic approaches based on high-resolution mass spectrometry; 2) to investigate specific lipidomic biomarkers in IOPD and LOPD patients.
Methodology: Plasma samples from untreated IOPD patients (n=12), LOPD (n=12), and healthy controls (adults: n=20; pediatric: n=13) were processed using a liquid-liquid extraction and analyzed with a quadrupole time-of-flight mass spectrometer coupled to ultra-performance liquid chromatography. Data were treated using Progenesis QI (Waters Corp.), and the MetaboAnalyst online platform.
Results: Several lipid classes, including glycerolipids, sphingolipids, and glycerophospholipids showed altered profiles in IOPD and LOPD patients compared to controls. Moreover, different lipid profiles were obtained between IOPD and LOPD patients. Sphingolipids were more altered in IOPD patients compared to LOPD patients, which might be due to the IOPD neuropathic involvement. Several lysophosphocholines were elevated in LOPD patients but not in the IOPD group. Phosphocholines with even- and odd-chain fatty acids were downregulated in Pompe disease patients. Preliminary longitudinal ERT treatment trend results showed that several dysregulated lipids tended to return to baseline, but heterogeneity was observed.
Conclusion: This study reveals altered lipidome profiles in PD patients. Future perspectives involve the validation/quantitation of the targeted biomarkers.
Gaucher disease (GD) is the most common lysosomal storage disorder and results from biallelic pathogenic variants in the GBA gene, which lead to deficient activity of the lysosomal enzyme, glucocerebrosidase. GD is classified into three subtypes based on the extent of central nervous system involvement. Although gallbladder disease has not traditionally been considered a feature of GD, limited studies estimate the prevalence of cholelithiasis in type 1 GD to be 2045%, compared with 615% in the general population. Data on the prevalence of gallbladder disease in GD types 2 and 3 remain unavailable. The mechanisms underlying this increased risk are not well understood but may involve increased bile lithogenicity and disrupted cholesterol metabolism. We conducted a retrospective chart review to obtain a more accurate estimate of the prevalence of gallbladder disease, including cholelithiasis and cholecystectomy, in individuals with GD. A secondary objective was to assess associations between prevalence and demographic characteristics, GD subtype, and disease severity. To date, six individuals treated at The Hospital for Sick Children as well as 50 treated at University Health Network and Sinai Health System have provided consent. Prevalence and risk factors will be presented to better delineate pathophysiology and guide screening and management recommendations regarding this emerging aspect of the GD phenotype.
CRISPR-Cas systems coupled to AAV-based delivery has been used to drive the site-specific integration of therapeutic transgenes into genomic safe harbors in neonatal mice. For liver-based in vivo genome editing, targeting transgenes to the first intron of the albumin gene allows for the potent secretion of coagulation factors, as well as several lysosomal enzymes. While secreted proteins can impact distant tissues in mouse models of lysosomal storage diseases (LSDs), achieving sufficient delivery across the blood-brain barrier (BBB) remains a key challenge. To achieve this, we engineered CRISPR-Cas9 systems to target therapeutic transgenes to the last intron of the transferrin gene. This leads to the generation of transferrin fusion proteins with improved secretion and tissue uptake since they can enter the cell via both the M6PR and the ubiquitous transferrin receptor. We have selected genes involved in LSDs for proof-of concept studies because they represent a paradigm for enzyme replacement therapies (ERTs). In mouse models of MPS1 and Pompe diseases, this approach led to stable and durable expression of fusion proteins in the serum, liver, heart, muscles, and brain. Compared to albumin-based strategies targeting intron 1, transferrin-targeted fusion proteins showed superior secretion and bioavailability. The lasting treatments prevented build-up of glycosaminoglycans (GAGs) in various organs and the loss of motor function in these mice models. The long-term objective of this project is to develop and test if generic liver-based enzyme replacement can cure or ameliorate the symptoms associated with rare and severe monogenic diseases affecting children.
Mucopolysaccharidoses (MPS) are lysosomal storage disorders caused by deficiencies in enzymes involved in glycosaminoglycan degradation. Although the primary enzymatic defects are well defined, their broader metabolic consequences remain incompletely characterized. Here, we used untargeted metabolomics and proteomics to identify molecular alterations in complementary mouse and Caenorhabditis elegans models of MPS. In an iduronate-2-sulfatase knockout mouse model of MPS II, liver samples from wild-type, untreated MPS II, and enzyme replacement therapy (ERT) treated animals were analyzed by LC-HRMS/MS. A total of 383 metabolites and 3,395 proteins were identified, with 96 metabolites and 220 proteins significantly altered between wild-type and MPS II mice. Major changes involved lysosomal proteins, acylcarnitine and lipid metabolism as well as redox-related metabolites.
To determine whether the molecular alterations observed in MPS II are also present in other MPS subtypes, complementary disease models were investigated. For this purpose, the same multi-omics strategy was extended to C. elegans models representing distinct MPS enzyme deficiencies. These datasets will be compared to determine which molecular alterations are shared across MPS models and which are specific to individual enzyme deficiencies. This multiomics strategy applied to two distinct models provides a large view of molecular changes associated with lysosomal dysfunction.
All sessions take place in salle de bal de Montréal 1-2-3
The classical phenotype of Fabry disease (FD) is associated with a reduced life expectancy. Over time the phenotypic spectrum of FD has widened, with an increase in the proportion of patients carrying GLA-variants associated with a milder, non-classical/late-onset form of the disease. Mortality associated with these variants hasnt been studied extensively. Simultaneously, life expectancy in the general population has increased. This complicates a comparison of survival between current- and historic cohorts and thus limits the analysis of impact of FD treatments such as enzyme replacement therapy.
We performed standardized mortality rate (SMR) analysis to study mortality associated with different phenotypes and individual GLA-variants in a collaboration between the Amsterdam UMC and Canadian Fabry Disease Initiative registry. The study involves a large cohort of 1042 FD patients and an additional 232 historic obligate carriers with data on mortality going back >150 years. SMR analysis, in contrast to Kaplan-Meier survival analysis, allows for correction of differences in life expectancy based on birth decade.
Mortality is clearly increased in males with classical GLA-variants (SMR 3.9, 95% CI 3.1-4.9) and to a lesser extent in women with these variants (SMR 1.6, 95% CI 1.3-2.0). Mortality isnt increased in patients with non-classical/late onset GLA-variants. Furthermore, the increased mortality correlates with untreated lysoGb3 levels, underscoring the predictive value of plasma lysoGb3 as a biomarker. Analysis of SMR trends over time showed no changes in SMR with the introduction of Fabry specific therapies such as ERT. SMR analysis can be used to assess the effect of treatment.
Male hemizygotes with classic Fabry disease (FD) are considered to be at high risk for complications with a marked reduction in life expectancy. As a result, treatment start is recommended by age 15 years in all classic males. There has been interest in applying this approach to females with classic FD. To determine whether a such recommendation for early treatment would be appropriate we studied treatment prevalence in women with severe classic FD in the CFDI registry. In the CFDI, 93 females had A143P GLA variant associated with severe classic phenotype. 70 were offered treatment under national guidelines. Enzyme replacement therapy (ERT) was the sole treatment as this variant is not enhanceable to chaperone. 62 patients mean age 61.014.8 years received ERT as 8 refused. Indications for ERT were mainly cardiac (62%). There were 11 deaths, mostly due to FD and 2 were lost to follow up (LTFU). The 31 untreated patients, mean age 39.719.5 years, had 2 FD deaths and 9 were LTFU. Fabry associated clinical event rates were higher in the ERT group than in the untreated patients, 66.7 vs 27.1/1,000 patient days. In conclusion, the untreated patients were younger, 25% had refused ERT and 29% were lost to follow up. While some of the untreated patients may need ERT as they age, it appears that not all women with classic FD will need or want ERT. Reasons for LTFU should be identified and minimized as this contributes to lower treatment rates in women with classic FD.
Heart Failure in Fabry Disease
Dr. Michael West, Halifax, NS
LAL Deficiency Cases
Dr. Régen Drouin, Laval, PQ
Posters are on display in salle de bal de Montréal 1-2-3
Background: Mucopolysaccharidosis type III is a spectrum (MPS IIIA-to-IIID) of rare neurodegenerative lysosomal storage disorders caused by genetic defects in heparan sulfate (HS) degradation, leading to its progressive accumulation. In the neurons, HS storage is associated with secondary accumulation of neurotoxic gangliosides and misfolded protein aggregates, including -amyloid, ultimately resulting in progressive neurodegeneration. To date, there is no clinically approved treatment for any type of MPS III. Although hematopoietic stem/progenitor cell (HSPC) transplantation has shown partial neuroprotective effect in the mouse MPS III models, its therapeutic efficacy remained limited (PMID:38786099). Our previous study found that lysosomal storage HS induces abnormal upregulation of lysosomal protease cathepsin B (CTSB), which contributes to amyloidogenesis and neurodegeneration (PMID:40092638). This project aims to evaluate whether combining the rain-permeable CTSB inhibitor E64 (Aloxistatin) with HSPC transplantation can improve neuropathology in the MPS III type C mice expressing mutant HGSNAT enzyme.
Methods: Memory and cognitive functions were assessed using multiple behavioral assays, enzymatic activities of HGSNAT and other lysosomal enzymes were measured in various organs, and immunofluorescence analysis was performed to assess neuronal levels of neurotoxic gangliosides and proteins and neuroinflammation.
Results: MPS IIIC mice transplanted with WT HSPC and treated with E64 showed behavioral improvements compared with mice treated with HSPC transplantation alone. They also showed normalized levels of activated astrocytes and partially normalized levels of Gm2 gangliosides and misfolded Thioflavin-S-positive proteins aggregates in the cortex. Notably, mice treated with a combination E64/HSPC therapy revealed increased residual levels of HGSNAT enzymatic activity, indicating partial correction of the underlying defect. Transcriptomic enrichment analysis revealed that E64 ameliorated apoptosis-related pathways in the brains of MPS IIIC mice compared to these receiving HSPC transplantation alone.
Conclusion: Together, our findings indicate that secondary induction of CTSB contributes to pathophysiological changes in the CNS in MPSA IIIC mice suggesting that CTSB may become a novel pharmacological target for MPS III and similar LSDs. Considering that E64 has been previously approved for clinical trials, its tempting to speculate that combination of E64 treatment with HSPC transplantation may become a promising therapeutic strategy.
DHDDS-congenital disorder of glycosylation (DHDDS-CDG) is a rare genetic disorder caused by pathogenic variants in DHDDS, which encodes the catalytic subunit of cis-prenyltransferase, an enzyme required for dolichol biosynthesis and protein glycosylation. Protein glycosylation is essential for cellular and organelle homeostasis, including the endolysosomal system, which supports lipid trafficking and degradation. The recurrent R37H variant is associated with developmental delay and epileptic myoclonic encephalopathy. DhddsR37H+/- knock-in mice exhibit behavioral abnormalities, myoclonus, and reduced density and maturation of brain interneurons. To investigate disease mechanisms, we differentiated patient-derived iPSCs carrying the heterozygous R37H mutation into neural progenitor cells and cortical neurons. Immunocytochemistry revealed elevated NPC2, lysosomal accumulation of GM1, GM2, and GM3 gangliosides, and increased COX IV-positive mitochondria compared with healthy controls. AVP6 reduced COX IV and LAMP2 accumulation and restored synaptic protein deficits in DHDDS neurons. These findings indicate altered lysosomal lipid homeostasis in DHDDS-CDG. We hypothesize that disruption of DHDDS function and dolichol-dependent glycosylation leads to secondary abnormalities in lysosomal and intracellular homeostasis, resulting in lysosomal accumulation of gangliosides, impaired neuronal and synaptic function, and neurological phenotypes. To evaluate a potential therapy, DhddsR37H+/- knock-in mice and wild-type littermates received daily intranasal AVP6, a synthetic ACTH analog with neuroprotective and neurotrophic properties, for three months. Open Field, Novel Object Recognition, and Contextual Fear Conditioning assessed behavior, while EEG and pentylenetetrazole (PTZ)-induced seizures evaluated neuronal excitability. Untreated DhddsR37H+/- knock-in mice showed hyperactivity and impaired learning and memory. AVP6 treatment rescued cognitive deficits and restored the reduced seizure threshold observed in mutant mice.
Sanfilippo type D, or mucopolysaccharidosis IIID (MPSIIID), is a rare lysosomal storage disorder (LSD) responsible for progressive debilitating neurodegeneration in children. MPS IIID is the result of genetic mutations preventing the enzyme N-acetylglucosamine-6-sulfatase (GNS) from degrading heparan sulfate (HS), which accumulates to toxic levels in tissues, leading to inflammation and neuronal death. Symptoms include worsening developmental delay, intellectual deficiency, cognitive decline, resulting in premature death during adolescence. No therapeutic treatment exists. One young MPSIIID patient is being treated at the CHU Sainte-Justine. Our objective is to develop an ex vivo hematopoietic stem cells (HSPCs) gene therapy for MPSIIID in a preclinical murine study and a worlds first single-patient clinical trial. We hypothesize that a lentiviral gene therapy correcting the patients HSPCs in an autologous transplantation will produce progenitors able to cross the blood-brain barrier and secrete sustained levels of functional GNS in the brain, to prevent further neurodegeneration.
In the preclinical trial, we have (1) demonstrated the safety of the treatment through biochemical and histological analyses in humanized mice, as well as an insertion site analysis, (2) successfully characterized a novel MPSIIID GNS-deficient mouse model through behavior analysis, enzymatic assay, and protein expression, and (3) are evaluating its efficacy in GNS-deficient mice by measuring enzymatic activity, behavior, gliosis, neuroinflammation, neuronal death and survival.
The goal of the project is to obtain approval from Health Canada to treat our patient, in order to prevent neurological deterioration and premature death; and establish a framework and reproducible therapy for other neurological LSDs.
Mucopolysaccharidoses (MPS) are diseases caused by mutations in genes coding for lysosomal enzymes that are responsible for the degradation of glycosaminoglycans (GAGs). These enzymes enable regulation of GAGs, including heparan-sulfate proteoglycans (HSPGs), which are abundant in the nervous system and play diverse roles in neural development and function. Patients with MPS accumulate HSPGs in excess, which can lead to severe nervous system degeneration in neuronopathic forms of MPS (types I, II, III VII). However, how HS deregulation impacts cellular biology remains poorly understood. We are dissecting this question in vivo by elucidating the cellular and molecular consequences of abnormal HSPG accumulation in the nervous system, using C. elegans models. To mimic the accumulation seen in MPS diseases, we study animals with mutations for genes orthologous to those underlying human MPS syndromes, as well as animals bearing an overabundance of three main HSPGs found in C. elegans. We use models for both neurodegenerative and non-neurodegenerative types of MPS, with the hope to help identify targets for neuronal manifestations. We thus examine the cellular and molecular impact of these conditions on neurons, associated glial cells and the extracellular matrix surrounding them. We also study the dynamics of protein degradation and autophagy on a cellular level as well as a global scale through unbiased proteomic and metabolomic analyses. These efforts will contribute to identifying new research avenues for MPS therapeutical strategies.
An open-label extension (OLE; NCT04138277) of the PROPEL study (NCT03729362) evaluated the long-term efficacy and safety of cipaglucosidase alfa plus miglustat (cipa+mig) in late-onset Pompe disease (LOPD). Outcomes are reported for enzyme replacement therapy (ERT)-experienced patients (median ERT duration at PROPEL baseline 7.6 years; n=62) and ERT-nave patients (n=20) who received cipa+mig from PROPEL baseline through the OLE. Data are presented as mean (standard deviation SD) change from PROPEL baseline (CFBL) at year 5 for ERT-experienced patients and year 4.5 for ERT-nave patients. In ERT-experienced patients, mean (SD) CFBL in % predicted 6-minute walk distance (pp6MWD) and % predicted forced vital capacity (ppFVC) was +0.7% (7.76%) and 2.8% (8.55%), respectively. For ERT-nave patients, mean (SD) CFBL in pp6MWD and ppFVC was +10.5% (11.44%) and 4.4% (6.89%), respectively. Mean (SD) CFBL in Rasch-built Pompe-specific Activity total score, Patient-Reported Outcomes Measurement Information System (PROMIS) Physical Function and PROMIS Fatigue scores was 1.3 (4.34), 1.1 (11.79) and 0.5 (6.97), respectively, in ERT-experienced patients and +2.1 (4.66), +6.3 (12.90) and 4.8 (4.81), respectively, in ERT-nave patients. Treatment-related treatment-emergent adverse events (TEAEs) occurred in 42/82 (51.2%) patients, leading to study drug discontinuation in five patients. Of these, two patients discontinued cipa+mig because of serious treatment-related TEAEs (chills and hypoxia n=1, anaphylactoid reaction n=1). Following up to 5 years of cipa+mig, patients generally experienced durable improvements or stabilisation in motor function, pulmonary function and patient-reported outcomes, supporting long-term benefits for patients with LOPD, even in those with a median 7.6 years on ERT prior to enrolment in PROPEL.
Background: Newborn screening for Mucopolysaccharidosis type I enables early detection; however, false positives and weak genotypephenotype correlation remain major challenges. Variants of uncertain significance (VUS) often exhibit reduced enzyme activity, without causing severe MPS-I. Classifying IDUA VUS is critical to identify severe form-Hurler syndrome, that benefits from early intervention with hematopoietic stem cell transplantation or enzyme replacement therapy.
Objectives: We establish a platform to functionally characterize IDUA variants using MPS-I patient-derived fibroblasts and a HEK293 IDUA knockout model.
Methods: First, LC-MS/MS-based IDUA activity and NRE-DS biomarker measurements, together with targeted LC-HRMS metabolomics were performed to build a variant classification framework. Second, lysosomal localization and protein processing studies assess whether IDUA VUS result in misfolded proteins, improper maturation, or incorrect localization. Third, MPS-I functional cellular phenotypes, including glycosaminoglycan (GAG) accumulation, lysosomal dysfunction, ER stress, autophagy, and cell death, categorize VUS by severity to guide clinical decisions.
Results: MPS-I samples showed significantly reduced IDUA activity and elevated GAG biomarkers compared with NHF, with metabolic profiling of Hurler fibroblasts revealing alterations in GAG-associated metabolites, including decreased glucuronic acid and N-acetyl-D-glucosamine, alongside broader metabolic changes. Lysosomal-tagged HEK 293 IDUA KO and IDUA variant bearing cell lines have been established for rapid organelle immunopurification which showed disrupted IDUA processing and lysosomal localization, improving pathogenic allele prediction. Lysotracker differences indicate lysosomal dysfunction, and potential additional cellular phenotypes can also be studied.
Conclusions: This integrated model will improve understanding of how IDUA variants cause disease and supporting earlier diagnosis and treatment for MPS I.
Heparan sulfate, a glycosaminoglycan, is covalently attached to core proteins forming heparan sulfate proteoglycans (HSPGs) found on the cell surface. Following endocytosis, HSPGs are degraded within lysosomes by several enzymes, including -N-acetylglucosaminidase (NAGLU). Absence of NAGLU leads to Mucopolysaccharidosis III type B (Sanfilippo type B), a lysosomal storage disease, which inNaglu-/-mice was suggested to result in infertility. Since endocytosis is a feature of epididymal epithelial and immune cells, a plausible explanation could be excessive lysosomal accumulation in these cells in 7-month-old mice. EM analysis revealed a massive accumulation of pale lysosomes (LM-IHC/LAMP2+/cathepsin B+) of varying shapes and sizes in principal cells, unlike the small, dense, spherical lysosomes of wild type mice. Their presence profoundly affected the integrity of the ER, Golgi, mitochondria, and nucleus. The epithelium also demonstrated sporadic large, vacuolated cells identified as CD68+ and MOMA+ epididymal mononuclear phagocytes (eMPs), with a cytoplasm dominated by pale lysosomes, some gigantic. The abundance of lysosomes suggested ongoing phagocytic activity by eMPs to protect the epithelium despite extensive organelle disruption. A distinguishing feature of eMP lysosomal content was aggregates of small flattened membranous discs aligned parallel to one another, referred to as zebra bodies. These resembled mitochondrial cristae and were reactive for a mitochondrial marker, suggesting phagocytosis by eMPs of degenerating principal cells. Taken together, our data demonstrate that lysosome accumulation in epithelial cells and eMPs dramatically altered their structural appearance in the absence of NAGLU, impairing their role in maintaining an epithelial environment for sperm protection and maturation.
DHDDS-associated progressive myoclonic epilepsy is a rare inherited disorder caused by pathogenic variants in the DHDDS gene, encoding cis-PTase, a dolichol synthesising enzyme essential for protein glycosylation pathway. The diseases causes developmental and cognitive impairment, myoclonus, seizures, and movement deficits, for which effective treatments are lacking. To study disease mechanism and test potential treatments we generated four genetically engineered induced pluripotent cell lines (iPSCs) carrying several patient-derived pathogenic DHDDS variants. These cells are reprogrammed into induced Pluripotent Stem Cells (iPSCs), which are then induced in Neuronal Progenitor Cells (NPC) and after differentiated into cortical neurons. Immunocytochemical analyses confirmed differentiation into cortical neurons by assessing Nestin, PAX6, and BCL11B expression. Disease-associated phenotypes were investigated, focusing on lysosomal accumulation of sphingolipids, a reported pathological feature of DHDDS-associated disorders. We found that unlike control cells, the neurons expressing DHDDS variants showed lysosomal storage vacuoles containing sphingolipids, including GM1, GM2 and GM3 gangliosides. They also demonstrated significantly increased LAMP2 levels, suggesting impaired lysosomal function. In addition, inhibitory (VGAT and Gephyrin) and excitatory (VGLUT and PSD95) synaptic markers were reduced, consistent with alterations in synaptic function and neuronal homeostasis. We further tested whether miglustat, an inhibitor of glycosphingolipid biosynthesis, approved for treatment of patients with Gaucher disease and Niemann-Pick disease type C (NPC), could reduce ganglioside storage. Our preliminary results demonstrate that GM1 accumulation was reduced in cells treated with miglustat during the neuronal differentiation phase, suggesting that this compound may partially alleviate cellular abnormalities observed in untreated neuronal models. Overall, our study validates engineered induced cultured neurons as relevant models for studying DHDDS-associated disorders. These models may provide valuable insight into cellular disease mechanisms and help to determine whether miglustat or other repurposed drugs effective in lysosomal sphingolipidoses can ameliorate disease-associated phenotypes.
Timely diagnosis of Lysosomal Storage Disorders (LSDs) remains challenging due to their rarity, symptom overlap with other conditions, and limited awareness among healthcare providers. For many LSDs, genetic testing is the most efficient and reliable way to confirm diagnosis, enabling earlier intervention and improved outcomes. However, access to testing is often hindered by cost, provider unfamiliarity, and complex diagnostic pathways.
To address these barriers, we developed a no-cost, patient-centric genetic testing program in the United States that leveraged artificial intelligence (AI) to identify at-risk individuals and offered whole exome sequencing (WES).
Probably Genetics AI-based Symptom Assessment was trained on anonymized profiles from LSD patients with systemic and neurologic manifestations. The predictive algorithm screened self-reported phenotype data from consenting US-based individuals to identify those at high likelihood of having an LSD. Testing, result delivery, and genetic counseling were conducted remotely through an independent physician network.
From October 2023 through April 2026, 714 participants were offered no-cost genetic testing. 417 (58%) buccal swab samples were returned and sequenced via WES. Thirteen individuals (3.1%) screened positive for LSD variants. Six pathogenic variants, five likely pathogenic variants, and four variants of uncertain significance were identified across 12 LSD-associated genes, including IDUA, SGSH, GUSB, NAGLU, MANBA, GLB1, and TPP1. Participants reported over 900 treating physicians across diverse specialties.
This program demonstrates how AI-guided, no-cost genetic testing can expand access to rare disease diagnosis and help close longstanding diagnostic gaps while highlighting the need for physician education and genetic counseling for families identified as carriers.

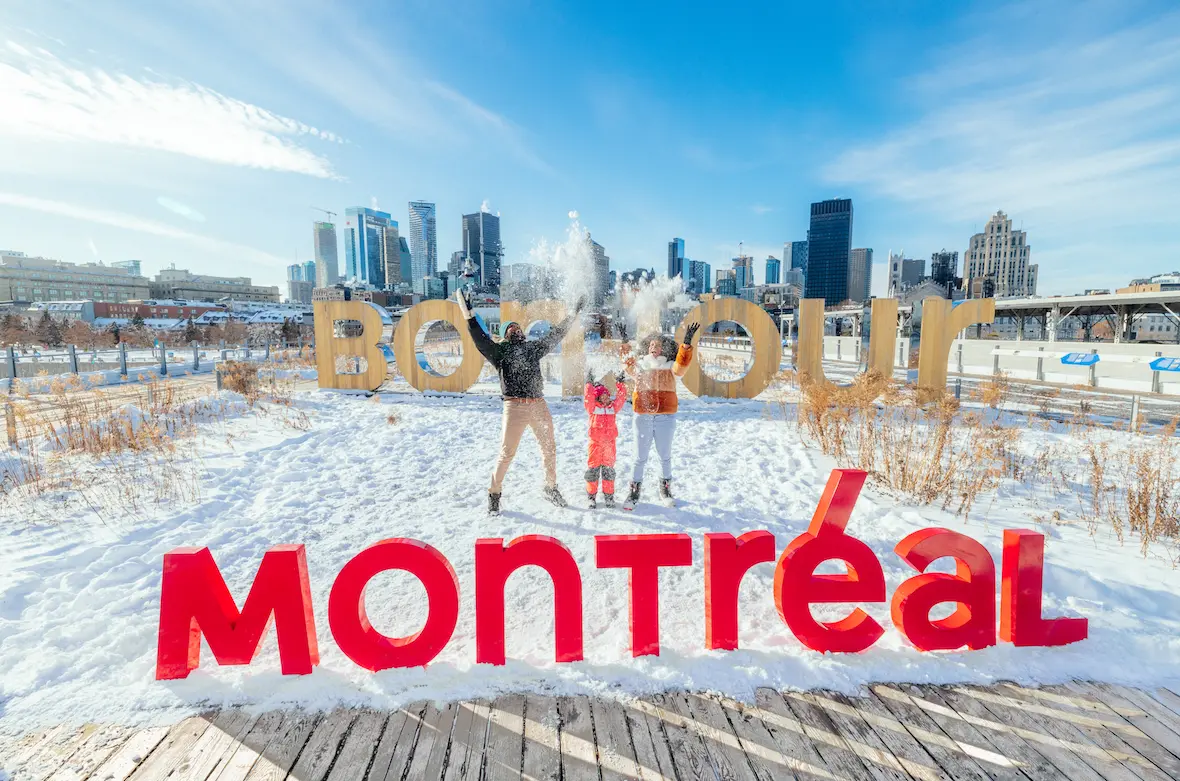
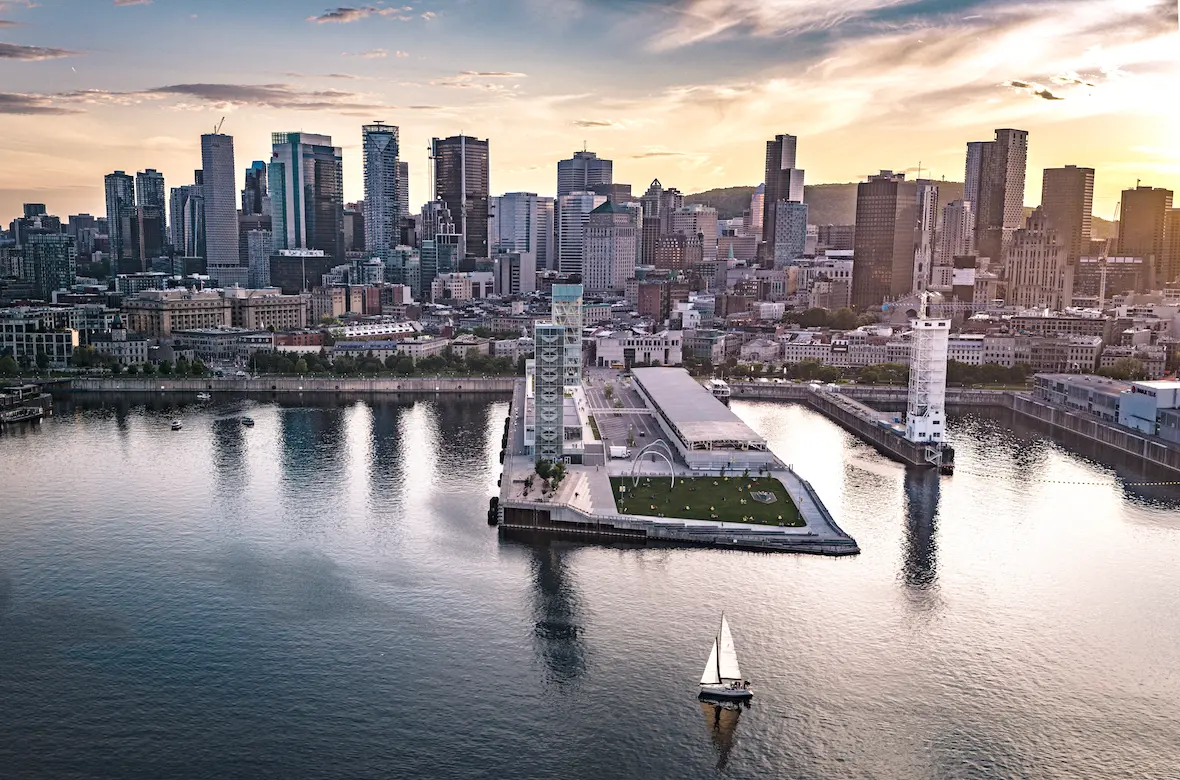
A striking union of European charm and North American attitude, Montréal seduces visitors with a harmonious pairing of the historic and the new, from exquisite architecture to fine dining. The world’s second largest francophone city after Paris, it merits the moniker ‘international’ city, a cosmopolitan centre with proud roots in the past that enthusiastically embraces the future. Its avant-garde spirit has not gone unnoticed: in 2006, Montréal was named a UNESCO City of Design. Please plan to take in all that Montréal has to offer.
Travelling to Montréal
Easy to get to from anywhere, Montréal, Montréal is connected to the world.
The newly revamped Montréal-Trudeau airport (YUL) offers direct flights to over 150 global destinations, along with more than 40 direct flights from cities throughout Canada.
Early bird registration ended on 29 October, 2026
All fees are stated in Canadian dollars (CAD).
*Upon arrival at the Symposium, students may be asked for ID verifying student status.
The deadline for bursary applications has passed and applicants have been notified.
Credit Card


Cheque
Please make cheques payable to CSLD and mail to:
CSLD c/o Agenda Managers
2979 Oxford Street, Halifax, Nova Scotia
Canada B3L 2W3
Wire Transfer
For details, please contact the CSLD Secretariat at .
All bank charges will be paid by the participant.
Cancellations received in writing prior to 15 October, 2026 will be charged a $25 administrative fee. Cancellations received after this date will not be refunded. Substitutions will be accepted with a request received in writing from the registered participant. Cancellations and substitutions should be forwarded to the CSLD Secretariat at .
Industry attendance at the Fourth Annual Canadian Symposium on Lysosomal Diseases (CSLD) is restricted to representatives of companies who sponsor the Symposium. Please contact Dr. Michael West to discuss sponsorship opportunities for the event and attendance.
The Fourth Annual Canadian Symposium on Lysosomal Diseases (CSLD) will be held at the Hotel Bonaventure Montréal, Montréal, Québec, 13 – 14 November, 2026. The Symposium will be held over 1-1/2 days with close to 100 clinicians, scientists, nurses, and genetic counsellors as well as trainees in attendance.
The Symposium offers leading experts in the field of lysosomal disease from Canada and abroad the opportunity to present and discuss current trends; learn from each other and subject matter experts; share best practices; and preview the latest and more relevant technologies and resources. Trainees will be encouraged to submit their research for poster and platform presentations. It will be in-person to allow for the interaction critical for a scientific Symposium.
CSLD 2026 offers an unprecedented opportunity to advocate for your organization by partnering with us. The investment impact for your organization includes, but is not limited to:
All fees are stated in Canadian dollars (CAD).
To confirm your commitment or learn more about opportunities to support CSLD 2026, please contact or .


Lysosomal storage diseases (LSDs) are a group of rare inherited metabolic disorders characterized by the dysfunction of lysosomes, cellular organelles responsible for breaking down various substances. In LSDs, there is a deficiency of specific enzymes within the lysosomes, leading to the accumulation of undigested materials inside the cells. This accumulation can cause damage to tissues and organs throughout the body.
There are more than 50 different types of LSDs, each associated with a specific enzyme deficiency and a unique set of symptoms. Examples of LSDs include Gaucher disease, Fabry disease, Niemann-Pick disease, and Pompe disease.
The Canadian Symposium on Lysosomal Diseases (CSLD Symposium) is an event focused on bringing together experts, researchers, clinicians, genetic counsellors, and other stakeholders to discuss the latest advancements, research findings, and treatment options related to lysosomal storage diseases. Attending the Canadian Symposium on Lysosomal Diseases can offer several benefits, including:
Education: Participants can gain insights into the current state of research and treatment options for lysosomal storage diseases. Experts in the field may present new findings, methodologies, and approaches to managing these rare disorders.
Networking: The symposium provides an opportunity for professionals and individuals involved in the field of lysosomal storage diseases to connect with one another. Networking can foster collaboration, information sharing, and the development of new research initiatives.
Advocacy and Awareness: The symposium may include discussions on advocacy efforts and raising awareness about lysosomal storage diseases. Increased awareness is crucial for fostering research, securing funding, and improving the overall understanding of these rare disorders.
Professional Development: Healthcare professionals, researchers, scientists, and other stakeholders attending the CSLD will advance their professional development, grow their knowledge and create collaborations that may be applied in their work to enhance patient care and outcomes.
If you are involved in the field of lysosomal storage diseases or have a personal or professional interest in these disorders, attending the CSLD Symposium could be a valuable opportunity to stay informed, connect with others in the field, and contribute to the ongoing efforts to improve the lives of individuals affected by LSDs.